


Clinical Characteristics
Ocular Features
Ectopia lentis as an isolated finding has been known for many years although early reports did not rule out features of the Marfan syndrome (154700). In more recent reports clinical evidence of the Marfan syndrome has been absent in a number of families and there seems little doubt that mutations in the FBN1 can be responsible for isolated ectopia lentis. Iridodenesis may be noted at birth but the dislocated lens may not be diagnosed until late adulthood in mild cases. Vision can be normal but nystagmus and strabismus have been noted in other patients. The lenses may be dislocated superiorly and may contain opacities. Areas of missing zonules have been observed in some patients while others have posterior synechiae.
Systemic Features
Related family members have been observed to have polydactyly and craniosynostosis but without dislocated lenses. It is important to rule out skeletal and cardiac manifestations of the Marfan syndrome because of the prognostic implications.
Genetics
Inheritance
This is an autosomal dominant disorder attributed to mutations in FBN1 (15q21), the same gene that is mutant in the Marfan syndrome (154700). The dislocated lenses may represent variable expressivity or simply allelism. The latter seems more likely in view of the fact that numerous thoroughly studied individuals have not had the skeletal or cardiovascular signs of the Marfan syndrome (154700). However, the revised Ghent nosology now suggests that all patients with the FBN1 mutation and ectopia lentis be designated to have the Marfan syndrome when aortic dilation/dissection is present as well. This should be extended to include all patients with FBN1 mutations and ectopia lentis plus at least one other phenotypic feature of the Marfan syndrome.
The same gene is mutant in the autosomal dominant form of the Weill-Marchesani 2 syndrome (608328) which is allelic to the Marfan syndrome.
There is also an autosomal recessive condition of isolated ectopia lentis (225100) which results from homozygous nonsense mutations in ADAMTSL4 (225100). A patient with craniosynostosis and ectopia lentis has been reported in which there was a homozygous 20 bp deletion in the same gene.
Pedigree
Autosomal dominant
Autosomal dominant disorders require only one mutation for the disease to be expressed. Since an affected parent has two chromosomes, only one of which has the mutant gene, parents can expect that half (50%) of their children will receive that one and inherit the disease. It is common for individuals that inherit the mutation, however, to not have evidence of the disease (nonpenetrance).
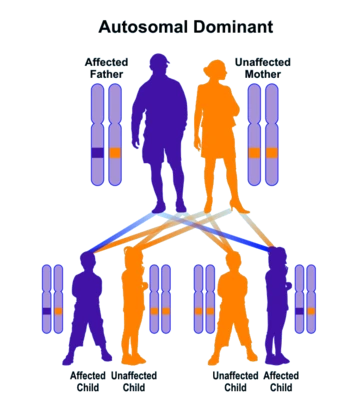
Autosomal dominant inheritance leads to a vertical pattern of transmission